What Is a Medical Device? Navigating FDA vs. EU MDR Definitions
In the global health technology ecosystem, the term "medical device" carries significant legal weight. It determines whether a product requires rigorous pre-market approval, undergoes clinical trials, or can be launched as a low-risk wellness tool. Yet, the definition of a medical device is not universal. While the United States Food and Drug Administration (FDA) and the European Union’s Medical Device Regulation (EU MDR) share a common goal—protecting public health—their legal frameworks, classification criteria, and interpretations of "intended use" diverge in critical ways.
For manufacturers, developers, and investors, understanding these distinctions is not optional; it is a prerequisite for market entry. Misclassifying a product can lead to enforcement actions, costly recalls, or the inability to sell in key markets. This article breaks down the legal definitions, classification systems, and the pivotal role of "intended use" under both regimes, providing a clear roadmap for compliance.
The Core Definition: Intended Use is King
At the heart of both the FDA and EU MDR frameworks lies a single, defining concept: intended use. Regulatory bodies do not judge a product solely by its technical capabilities but by the claims made about its purpose.
The FDA Perspective
Under the Federal Food, Drug, and Cosmetic Act (FD&C Act), a medical device is defined as an instrument, apparatus, implement, machine, contrivance, implant, in vitro reagent, or other similar article that is:
- Recognized in the official National Formulary or the United States Pharmacopeia.
- Intended for use in the diagnosis, cure, mitigation, treatment, or prevention of disease in man or other animals.
- Intended to affect the structure or any function of the body of man or other animals, and which does not achieve its primary intended purposes through chemical action within or on the body.
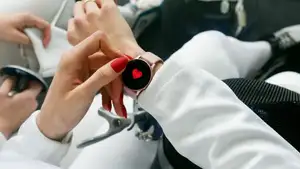
Crucially, the FDA emphasizes that the intended use is determined by the claims made by the manufacturer in labeling, advertising, or promotional materials. If a smartwatch claims to "track steps," it is a wellness product. If it claims to "detect atrial fibrillation," it instantly becomes a Class II medical device, regardless of the hardware's identical nature.
The EU MDR Perspective
The EU Medical Device Regulation (Regulation (EU) 2017/745) offers a similarly broad but distinct definition. A medical device is any instrument, apparatus, appliance, software, implant, reagent, material, or other article intended by the manufacturer to be used, alone or in combination, for human beings for one or more of the following specific medical purposes:
- Diagnosis, prevention, monitoring, prediction, prognosis, treatment, or alleviation of disease.
- Diagnosis, monitoring, treatment, alleviation of, or compensation for, an injury or disability.
- Investigation, replacement, or modification of the anatomy or of a physiological or pathological process or state.
- Providing information by means of in vitro examination of specimens derived from the human body.
The EU MDR places a heavier emphasis on the manufacturer’s intention as declared in the technical documentation and labeling. Unlike the FDA, which historically focused on "chemical action" to distinguish drugs from devices, the EU MDR explicitly includes software as a medical device (SaMD) and clarifies that products achieving their action via pharmacological, immunological, or metabolic means are generally excluded unless they assist in a device’s function.
Classification Systems: Risk-Based Tiers
Both jurisdictions classify devices based on risk, but the tiers and criteria differ significantly.
FDA Classification (US)
The FDA categorizes devices into three classes based on the level of control necessary to ensure safety and effectiveness:
- Class I (Low Risk): Subject to general controls (e.g., labeling, good manufacturing practices). Examples: Elastic bandages, manual stethoscopes, non-electric wheelchairs. Most are exempt from pre-market notification (510(k)).
- Class II (Moderate Risk): Require general controls plus special controls (e.g., performance standards, post-market surveillance). Most require a 510(k) pre-market notification to demonstrate substantial equivalence to a legally marketed predicate device. Examples: Infusion pumps, powered wheelchairs, pregnancy test kits.
- Class III (High Risk): Devices that support or sustain human life, are of substantial importance in preventing impairment of health, or present a potential unreasonable risk of illness or injury. These require Premarket Approval (PMA), the most stringent pathway, often necessitating clinical trial data. Examples: Implantable pacemakers, heart valves, HIV diagnostic tests.
EU MDR Classification (Europe)
The EU MDR uses a four-class system (I, IIa, IIb, III) governed by 22 classification rules based on duration of use, invasiveness, and the part of the body affected:
- Class I (Low Risk): Non-sterile, non-measuring devices (e.g., surgical masks, crutches). Self-declaration is permitted; no notified body involvement is required unless sterile or measuring.
- Class IIa (Low-to-Moderate Risk): Short-term invasive devices or devices for monitoring physiological processes (e.g., hearing aids, dental fillings). Requires assessment by a Notified Body.
- Class IIb (Moderate-to-High Risk): Long-term invasive devices, implantable devices, or those that administer/remove drugs (e.g., infusion pumps for high-risk drugs, ventilators). Requires Notified Body assessment.
- Class III (High Risk): High-risk implantable devices or those intended to diagnose/treat life-threatening conditions (e.g., breast implants, drug-eluting stents, HIV tests). Requires Notified Body assessment and often clinical investigation.
Key Difference: The EU MDR has expanded the scope of devices requiring Notified Body intervention. Many products that were Class I under the old Directive (MDD) are now Class IIa or IIb under the MDR, particularly those with software components or measuring functions.
The "Intended Use" Trap: Common Pitfalls
The most frequent regulatory misstep occurs when companies blur the line between wellness and medical claims.
- Software Claims: An app that tracks sleep patterns for "better rest" is wellness. The same app claiming to "diagnose insomnia" or "predict seizure risk" becomes a medical device.
- Algorithmic Diagnostics: AI algorithms analyzing retinal images for "screening" may be wellness, but if they claim to "detect diabetic retinopathy," they are Class II (FDA) or Class IIa/IIb (EU).
- Wellness Wearables: A fitness tracker measuring heart rate for "exercise intensity" is safe. If it markets itself as a "heart rhythm monitor" for detecting arrhythmias, it triggers medical device regulations.
Manufacturers must rigorously audit all marketing materials, user manuals, and software interfaces to ensure claims align with their intended regulatory pathway. A single phrase in a press release can redefine a product’s entire regulatory status.
Strategic Implications for Global Market Entry
Navigating these differences requires a dual-track strategy. A product designed for the US market may need significant adjustments to comply with the EU MDR, and vice versa.
- Documentation: The EU requires a comprehensive Technical File and a Declaration of Conformity, while the US focuses on the 510(k) or PMA submission.
- Timeline: The FDA 510(k) process typically takes 3–6 months, whereas EU Notified Body reviews can take 12–18 months due to backlog and increased scrutiny.
- Post-Market Surveillance: Both require vigilance, but the EU mandates a stricter Post-Market Clinical Follow-up (PMCF) and a Unique Device Identification (UDI) system integrated into the EUDAMED database.
Conclusion
Defining a medical device is the first and most critical step in the regulatory journey. Whether navigating the FDA’s risk-based classes or the EU MDR’s intricate rules, the mantra remains the same: intended use dictates regulation. By aligning product claims with the appropriate regulatory framework, manufacturers can avoid costly delays, ensure patient safety, and successfully bring innovative health technologies to market.
Frequently Asked Questions (FAQ)
Q: Can a product be a medical device in the EU but not in the US?
A: Yes. The EU MDR has a broader definition that includes certain software and products for contraception or abortion prevention, which may not be classified as medical devices under FDA rules if they don’t meet the "diagnosis/treatment" threshold. Conversely, some FDA Class I devices might face stricter scrutiny in the EU if they involve measuring functions.
Q: Does the FDA consider software a medical device?
A: Yes, if the software is intended for a medical purpose. The FDA regulates Software as a Medical Device (SaMD) under the same framework as hardware, classifying it based on risk and intended use.
Q: What is a "predicate device" in the FDA 510(k) process?
A: A predicate device is a legally marketed device to which a new device is compared to demonstrate substantial equivalence. It must be already cleared by the FDA for the same intended use.
Q: How does the EU MDR handle "wellness" products?
A: The EU MDR does not have a specific "wellness" category. Products that do not meet the medical purpose definition are not medical devices. However, the burden of proof is on the manufacturer to demonstrate that a product is not intended for medical use.
Q: What happens if I make a medical claim for a wellness product?
A: Regulators may reclassify the product as an unapproved medical device, leading to warning letters, product seizures, fines, or legal action. Marketing claims are the primary determinant of regulatory status.
Q: Where can I find the official FDA and EU MDR definitions?
A:
Disclaimer: This article is for informational purposes only and does not constitute legal or regulatory advice. Always consult with qualified regulatory professionals for specific guidance on your products.
Sources:
- U.S. Food and Drug Administration. (2023). Classification of Medical Devices. Retrieved from FDA.gov
- European Commission. (2017). Regulation (EU) 2017/745 on medical devices. EUR-Lex.
- U.S. Food and Drug Administration. (2021). Software as a Medical Device (SaMD). Retrieved from FDA.gov
- International Medical Device Regulators Forum (IMDRF). (2020). Definitions and Terminology.